Prospective discovery of LasB inhibitors by integrating generative design, free energy ranking, and experimental validation
Abstract
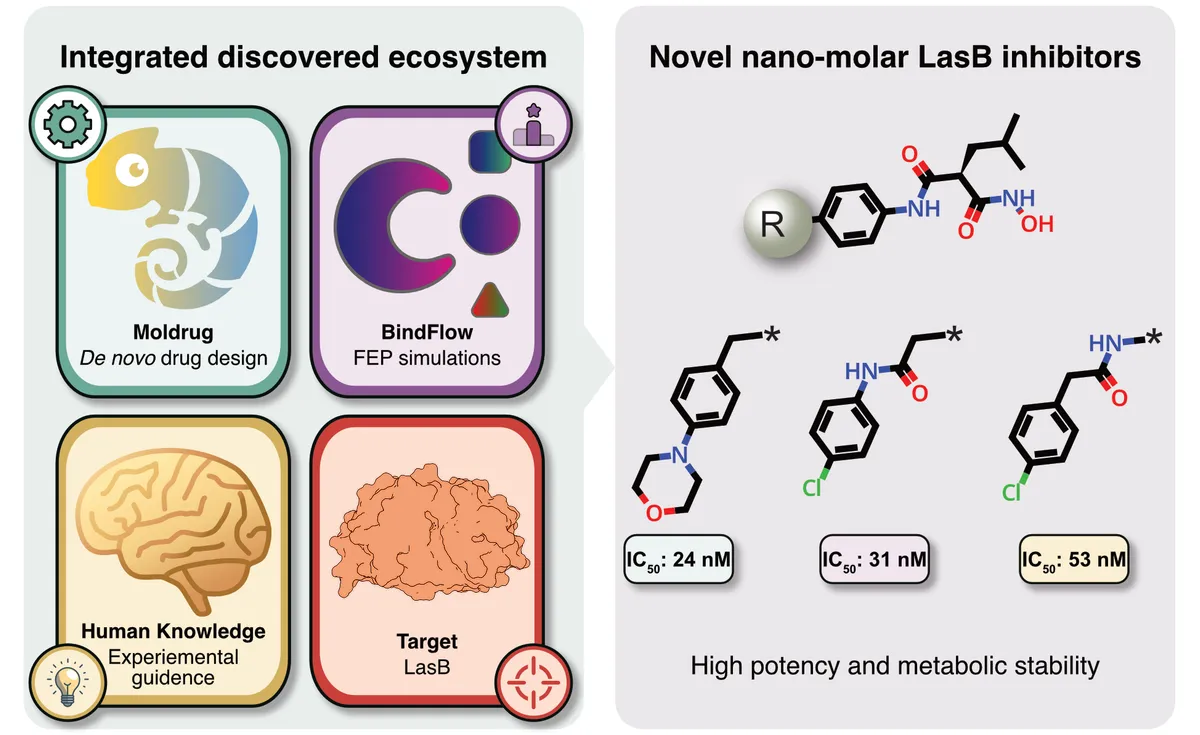
Identifying synthetically accessible, potent ligands from vast chemical space remains a major challenge in drug discovery, particularly because experimental synthesis and testing are costly and low throughput. Here, we present a fully open-source computational–experimental workflow that integrates fragment-based de novo molecular design with rigorous alchemical absolute binding free-energy calculations to enable physics-based prioritization before chemical synthesis. We apply this strategy by targeting the virulence-associated zinc metalloprotease LasB from Pseudomonas aeruginosa. From ~40,000 diverse compounds generated through multi-objective optimization, a small subset was prioritized and experimentally evaluated, leading to the discovery of three previously unreported nanomolar inhibitors (IC50 22–53 nM) with excellent metabolic and plasma stability. This work demonstrates a general and scalable framework for coupling generative molecular design with quantitative binding affinity prediction, improving decision-making and reducing the cost, time, and risk associated with early-stage drug discovery..
Cite
@article {leon2026lasb,
	author = {Mart{\'\i}nez Le{\'o}n, Alejandro and Vgenopoulou, Angeliki and J{\'e}z{\'e}quel, Gwena{\"e}lle and Kany, Andreas M and Hirsch, Anna KH and Hub, Jochen S},
	title = {Prospective discovery of LasB inhibitors by integrating generative design, free energy ranking, and experimental validation},
	elocation-id = {2026.10.12.758448},
	year = {2026},
	doi = {10.64898/2026.10.12.758448},
	publisher = {Cold Spring Harbor Laboratory},
	abstract = {Identifying synthetically accessible, potent ligands from vast chemical space remains a major challenge in drug discovery, particularly because experimental synthesis and testing are costly and low throughput. Here, we present a fully open-source computational{\textendash}experimental workflow that integrates fragment-based de novo molecular design with rigorous alchemical absolute binding free-energy calculations to enable physics-based prioritization before chemical synthesis. We apply this strategy by targeting the virulence-associated zinc metalloprotease LasB from Pseudomonas aeruginosa. From ~40,000 diverse compounds generated through multi-objective optimization, a small subset was prioritized and experimentally evaluated, leading to the discovery of three previously unreported nanomolar inhibitors (IC50 22{\textendash}53 nM) with excellent metabolic and plasma stability. This work demonstrates a general and scalable framework for coupling generative molecular design with quantitative binding affinity prediction, improving decision-making and reducing the cost, time, and risk associated with early-stage drug discovery.Competing Interest StatementThe authors have declared no competing interest.European Union, 860592, 101081463Deutsche Forschungsgemeinschaft, INST 256/539-1},
	URL = {https://www.biorxiv.org/content/early/2026/10/13/2026.10.12.758448},
	eprint = {https://www.biorxiv.org/content/early/2026/10/13/2026.10.12.758448.full.pdf},
	journal = {bioRxiv}
}